 4 Images
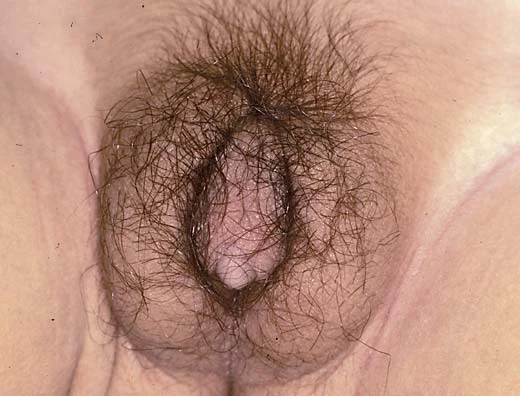
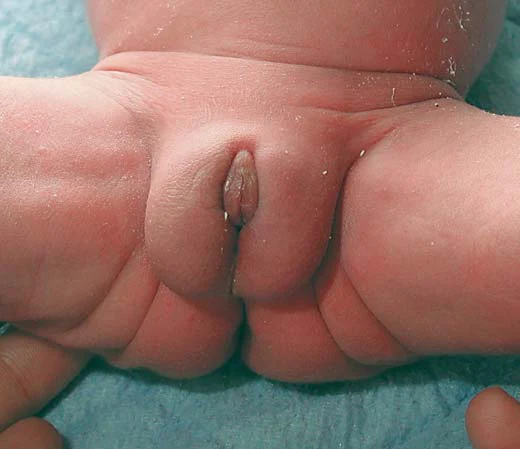
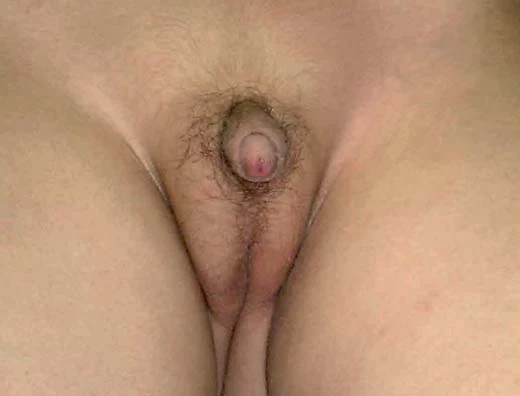
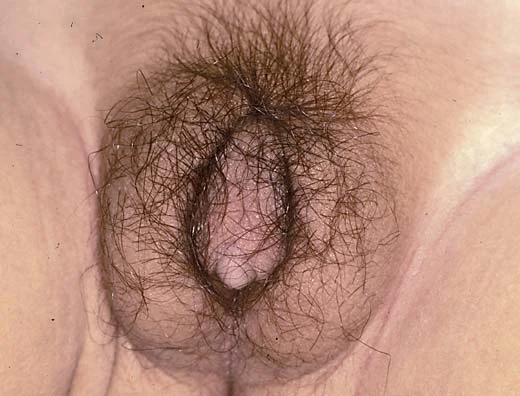
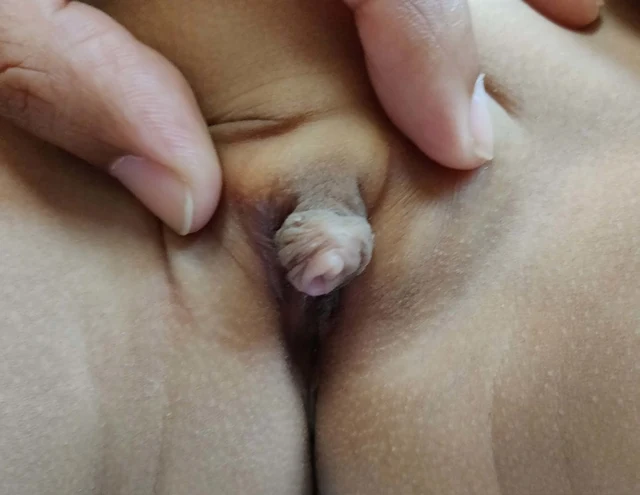
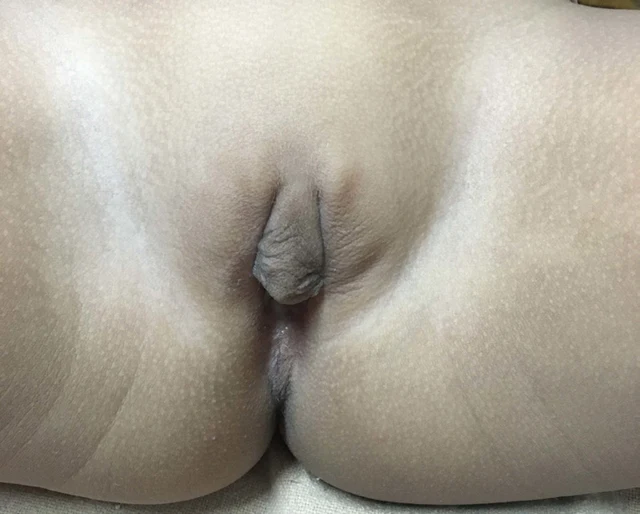
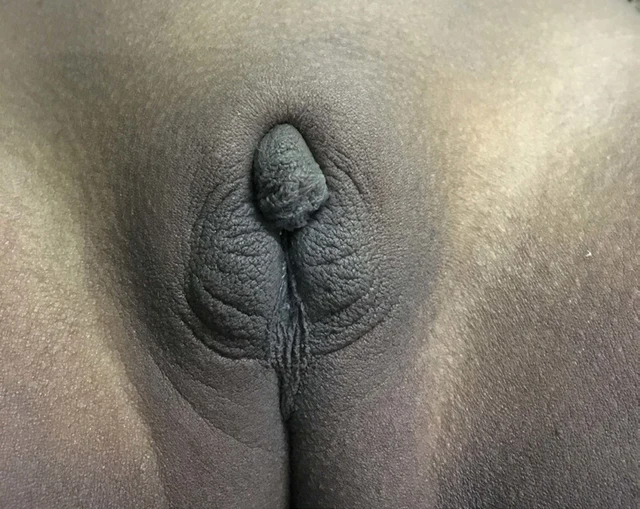
5α-reductase deficiency · 46,XY · DSD-C121
46,XY DSD (SRD type III / Prader-like classification) caused by 5α-reductase type 2 deficiency (SRD5A2 defect).
Browse cases and figures from the original collection. Search by condition, gene, karyotype or phenotype and follow the source publication.
280 Image cases571 Images
Current selection: 280 Image cases · 571 Images
Select an image to enlarge it. Original images retain the complete source file.
4 Images
46,XY DSD (SRD type III / Prader-like classification) caused by 5α-reductase type 2 deficiency (SRD5A2 defect).
 1 Images
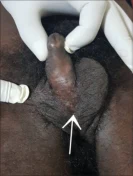
Author's diagnosis: 45,X0/46,XY mixed gonadal dysgenesis (MGD) with persistent Müllerian structures manifesting as symptomatic pelvic cysts.
 4 Images
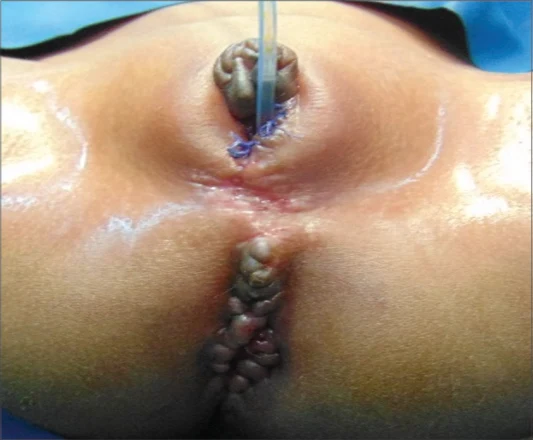
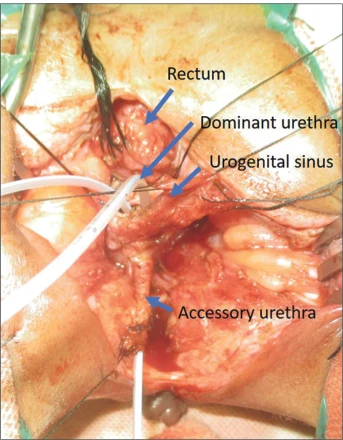
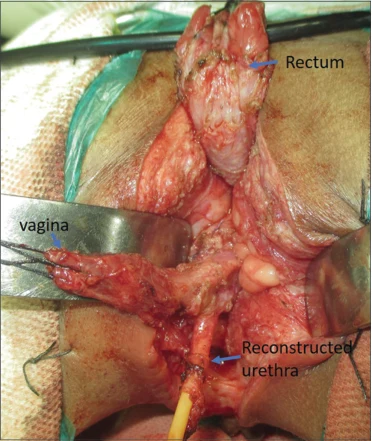
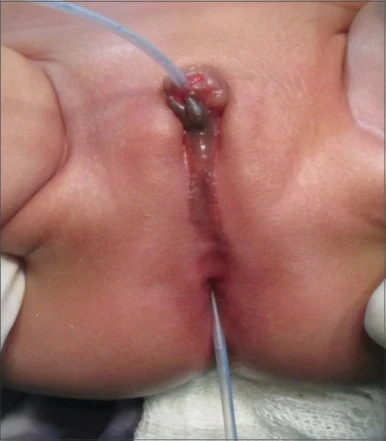
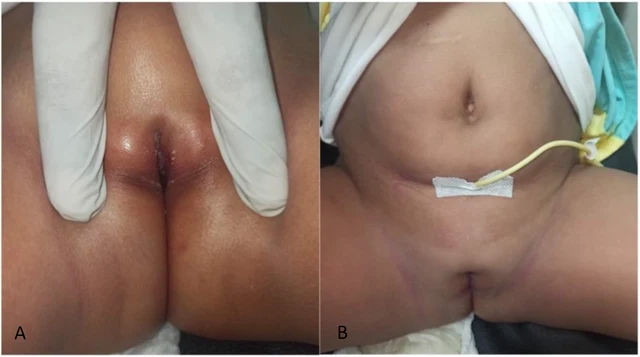
46,XX disorder of sex development with type-A posterior cloacal malformation and Y-type urethral duplication (rare variant).
 2 Images
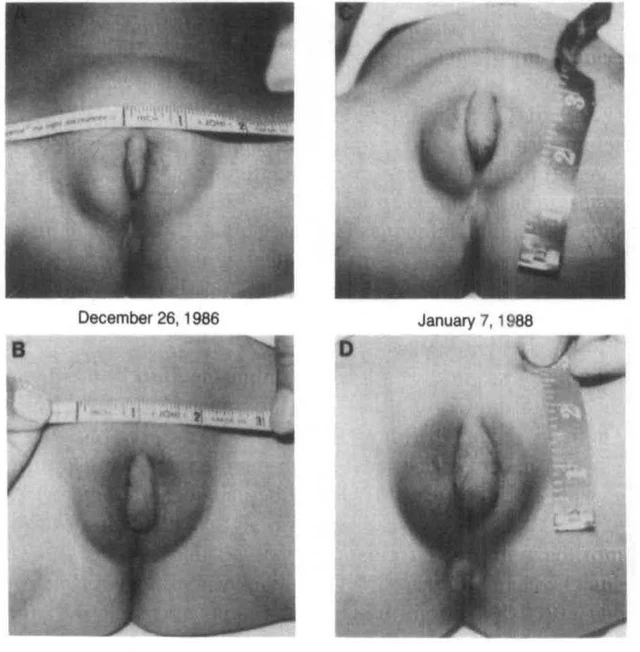
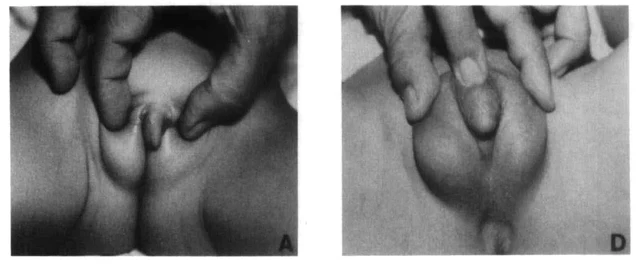
Androgen resistance syndrome — partial male pseudohermaphroditism due to a qualitative androgen receptor defect. This case is unique in showing a marked response to high-dose androgen therapy (overcoming receptor dysfunction).
 2 Images
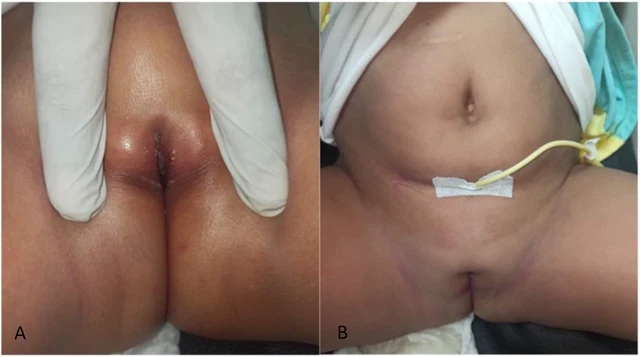
46,XX disorder of sex development with high confluence urogenital sinus (UGS). First report of traction vaginoplasty via the anterior sagittal transrectal approach (ASTRA).
 2 Images
46,XX disorder of sex development — aromatase deficiency (AD). Often misdiagnosed as CAH 21-OHD. AD features: non-progressive external virilization + maternal androgen excess + delayed bone age + hypergonadotropism.
 1 Images
46,XX disorder of sex development — aromatase deficiency, more severe phenotype (Prader stage 3). Sibling/series case with Case 1.
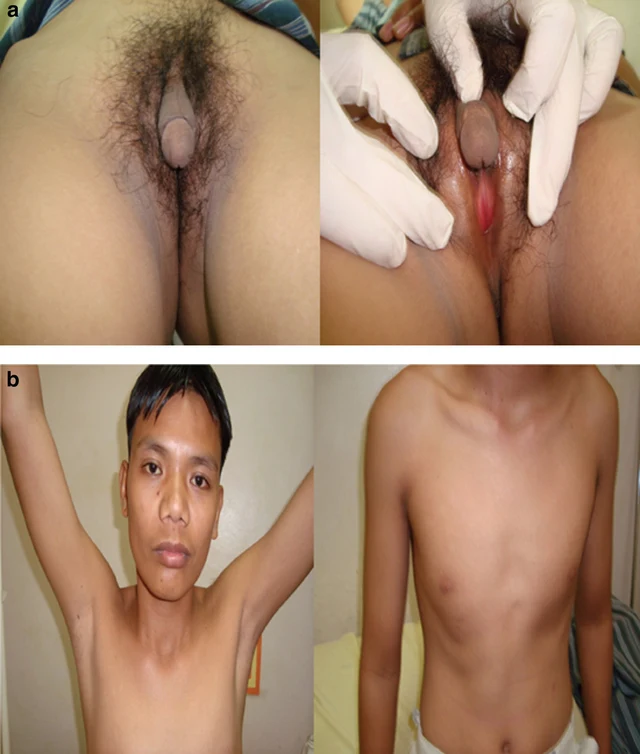 1 Images
46,XY gonadal dysgenesis (complete/incomplete intermediate state) — a 'late-diagnosis trap' in adulthood. Key lesson: the importance of early correct sex assignment and the profound psychosocial impact on DSD patients.
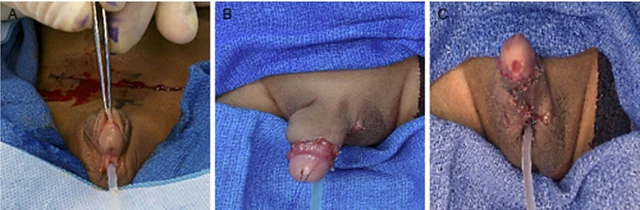 1 Images
Ovotesticular DSD with mosaic karyotype 45,X/46,X,idic(Y)(q11.23), with streak gonads and ambiguous genitalia.
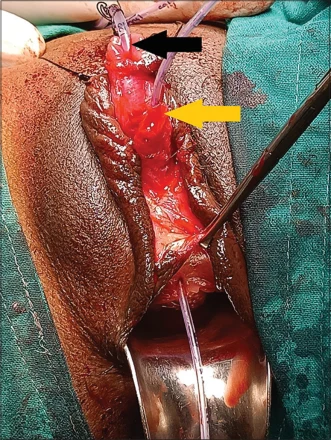 2 Images
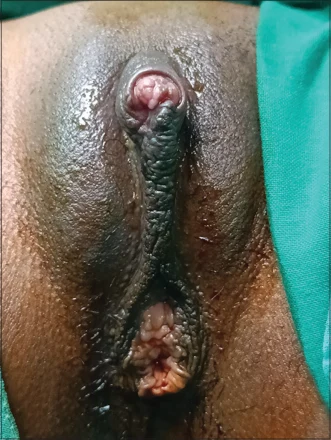
Female pseudohermaphroditism (likely 46,XX CAH) with urethral duplication, late-onset presentation in adulthood.
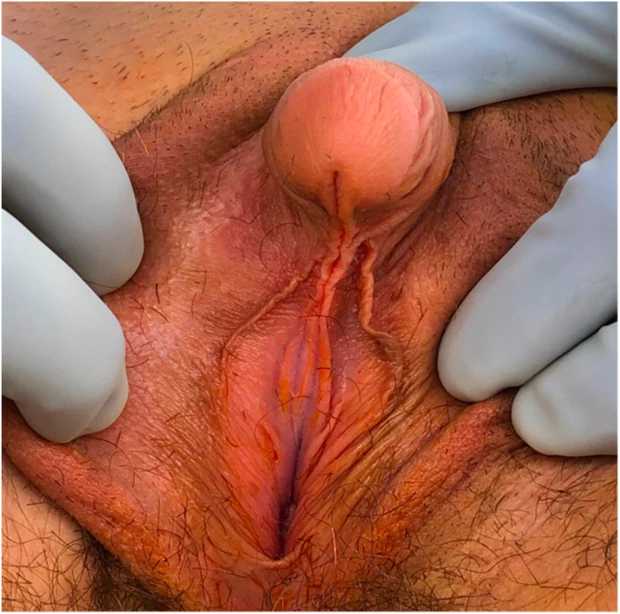 1 Images
46,XY DSD with severe hypospadias, severe chordee, and proximal urethral meatus.
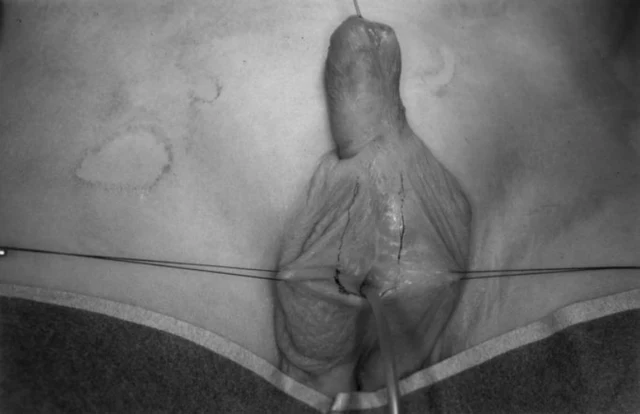 1 Images
Mixed gonadal dysgenesis (45,X/46,X,idic(Y) mosaic) with scrotal hypospadias and coarctation of the aorta.
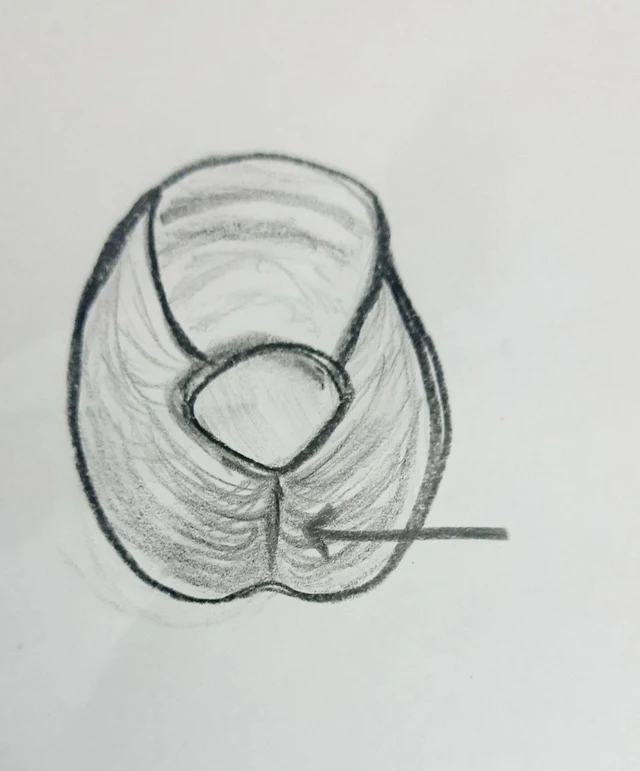 3 Images
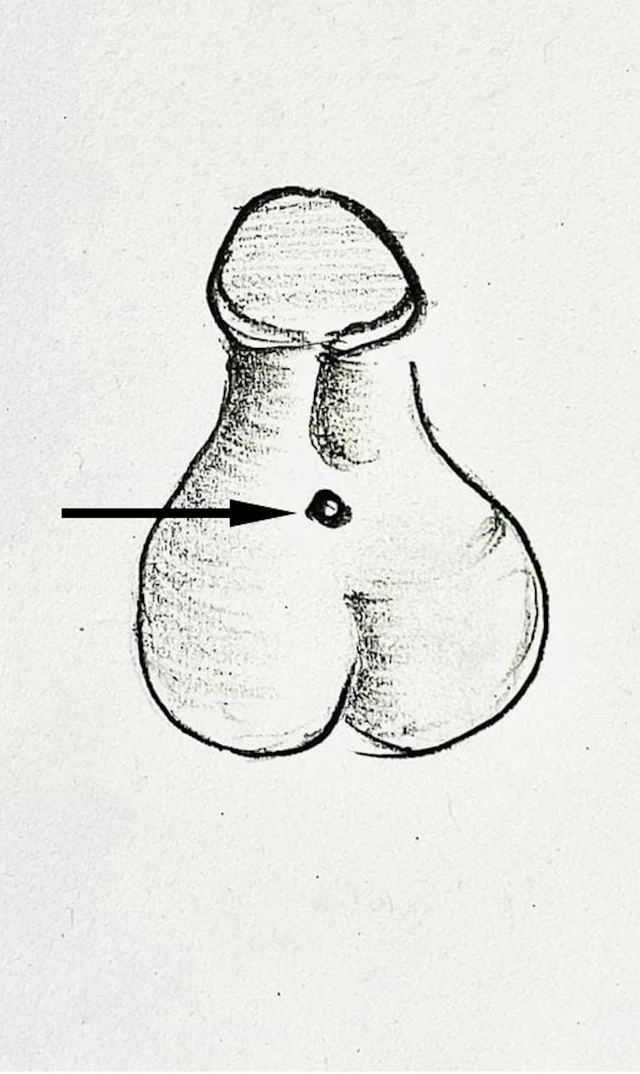
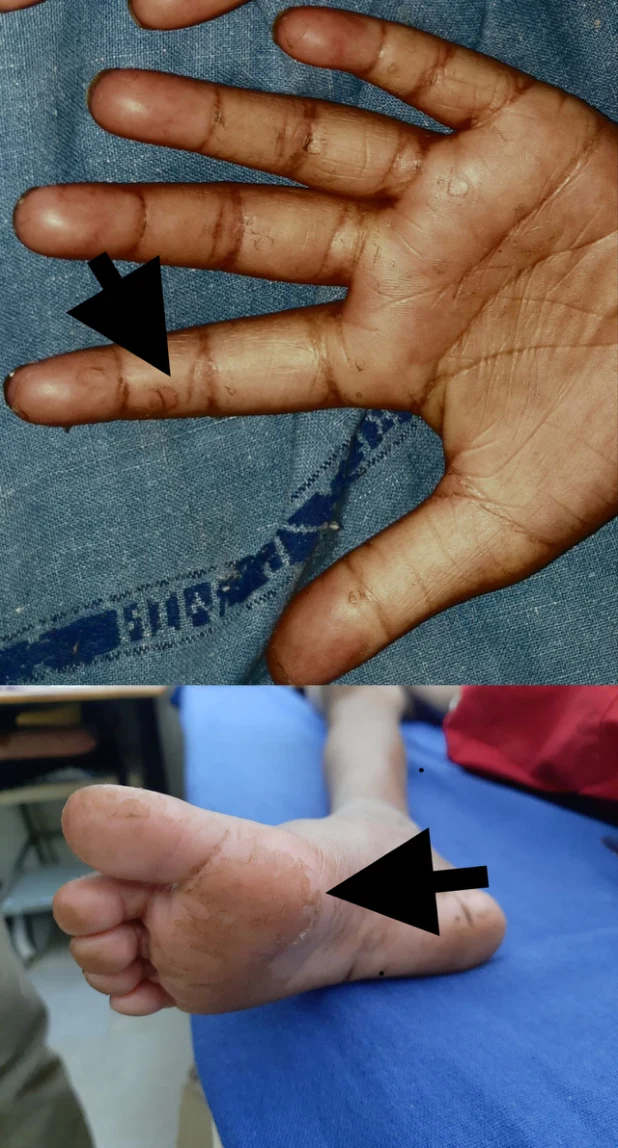
46,XX DSD with homozygous loss-of-function mutation in R-Spondin 1 (RSPO1), with cutaneous anomalies and virilized phenotype.
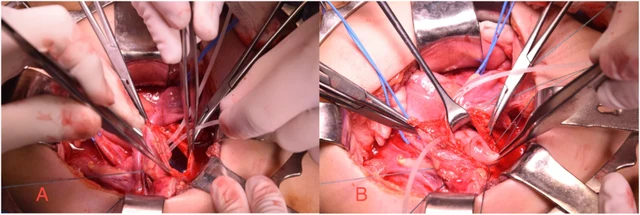 2 Images
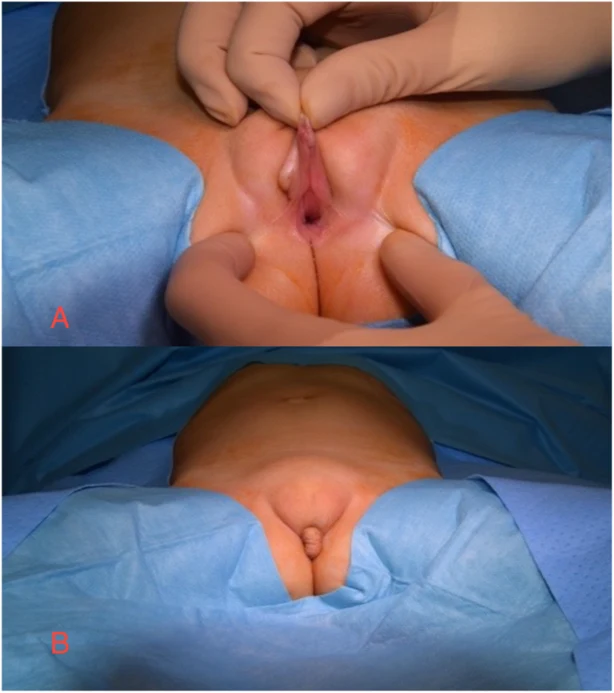
Complex urogenital malformation (unclassified) in a 3-year-old girl, with ambiguous genitalia and cloacal/urinary tract anomalies.
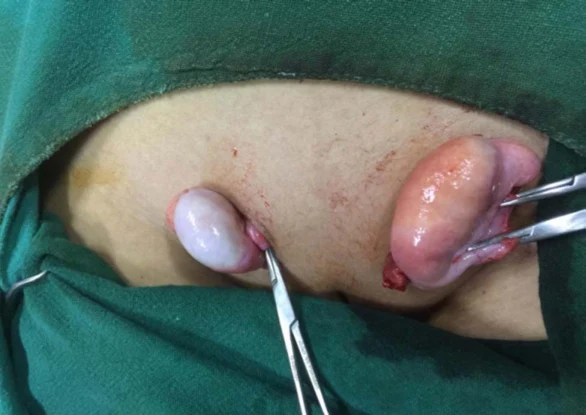 1 Images
Complete androgen insensitivity syndrome (CAIS), confirmed after inguinal surgery.
 3 Images
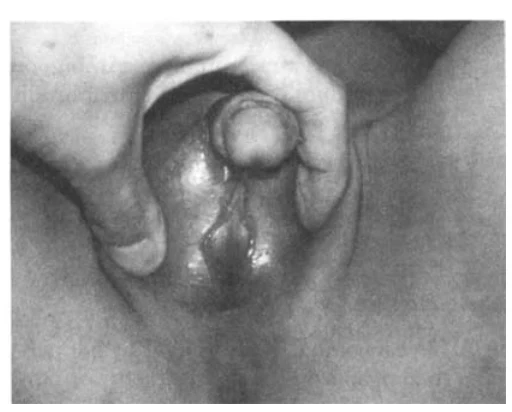
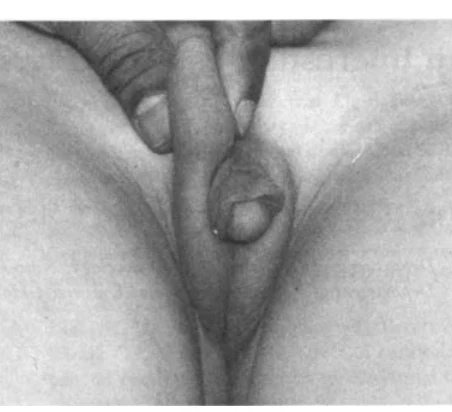
Intersex case: clitoromegaly with labioscrotal fusion (likely 46,XX CAH).
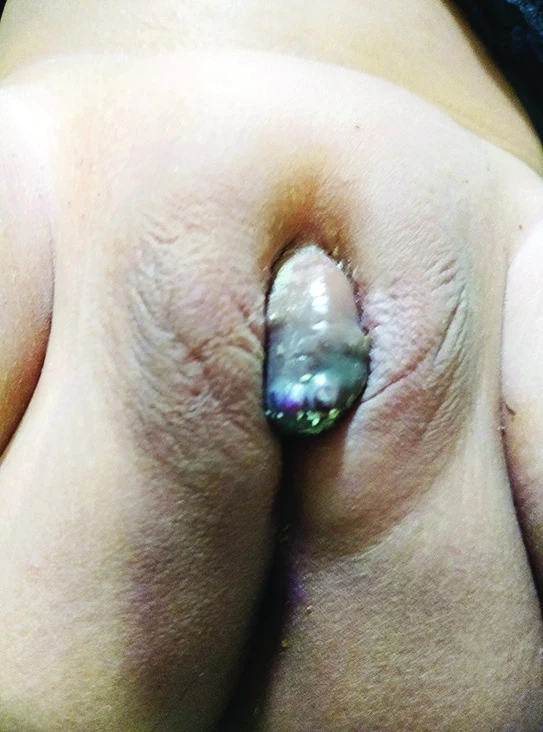 1 Images
Clitoromegaly in a girl (non-hormonal etiology, with perineal hemangioma).
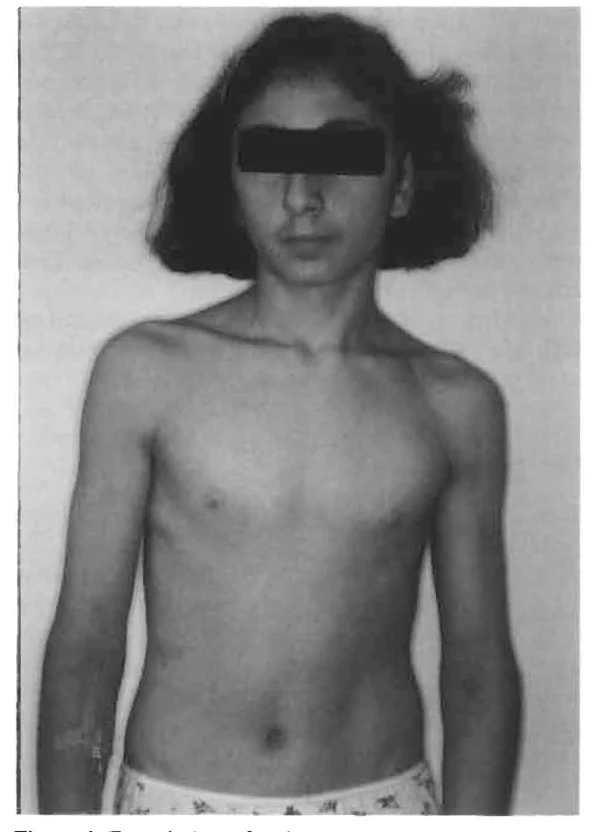 2 Images
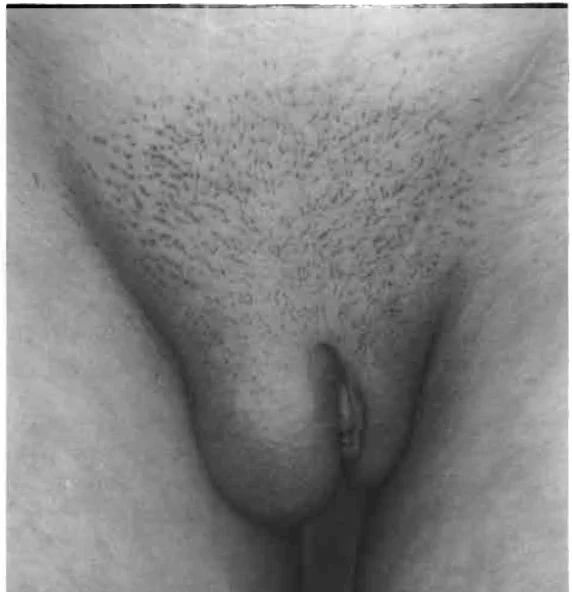
5α-reductase type 2 deficiency, adolescent female phenotype (46,XY).
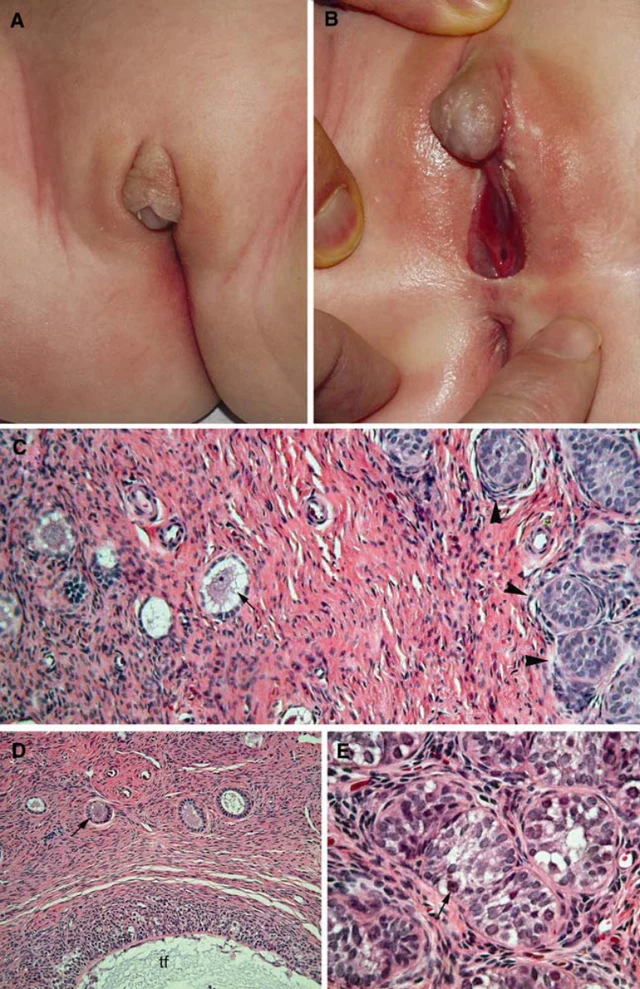 1 Images
True hermaphroditism (ovotesticular DSD), with an abnormal twinning pathogenesis.
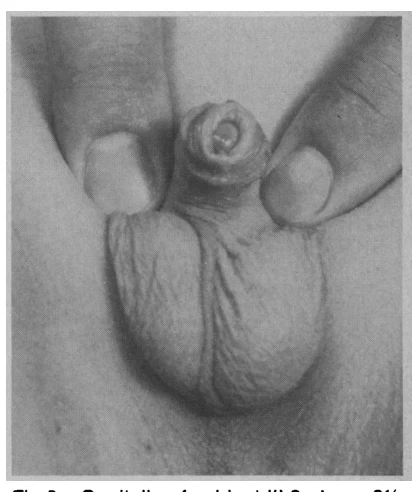 1 Images
Partial androgen insensitivity syndrome (PAIS) family, with highly variable phenotype.
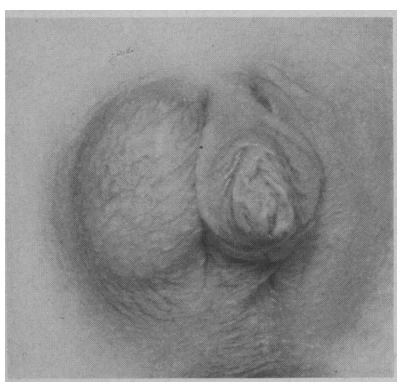 1 Images
Familial partial androgen insensitivity syndrome (PAIS) with highly variable phenotype.
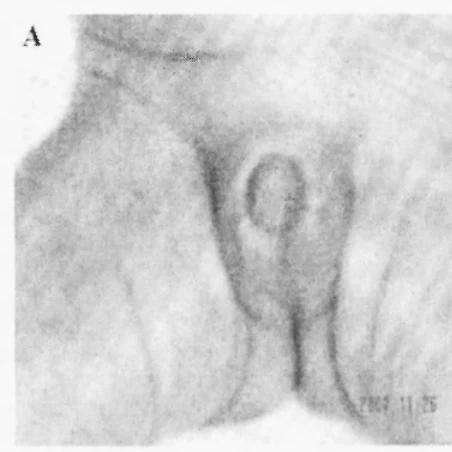 3 Images
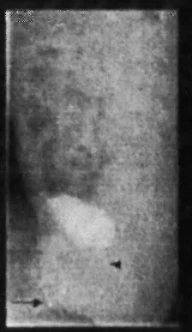
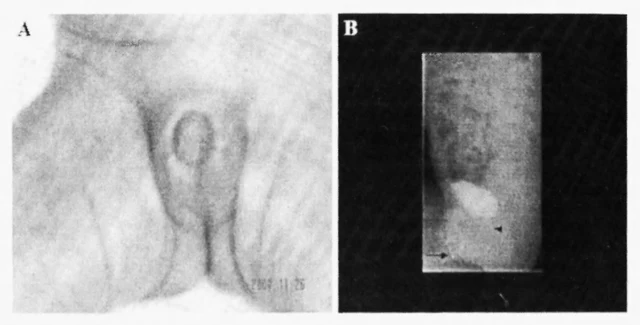
Sex reversal (46,XX CAH) caused by donor splice mutation in 11β-hydroxylase (CYP11B1).
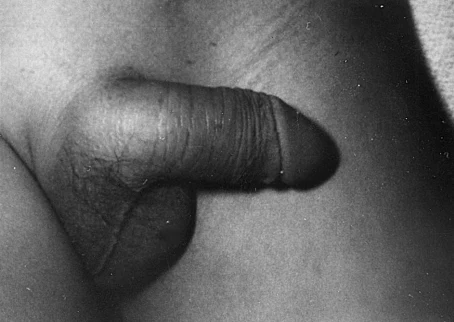 1 Images
Boy with deletion of the long arm of chromosome 10 accompanied by monorchidism.
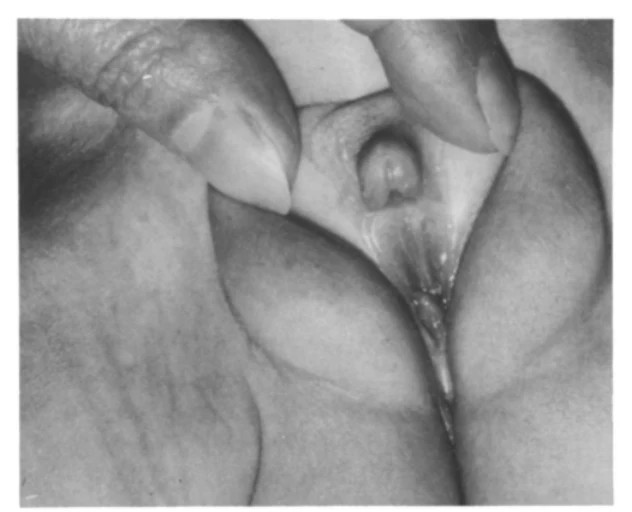 1 Images
Drash syndrome: partial androgen receptor defect with mixed gonadal dysgenesis (46,XY).