 1 Images
46,XX DSD · 46,XX · DSD-C098
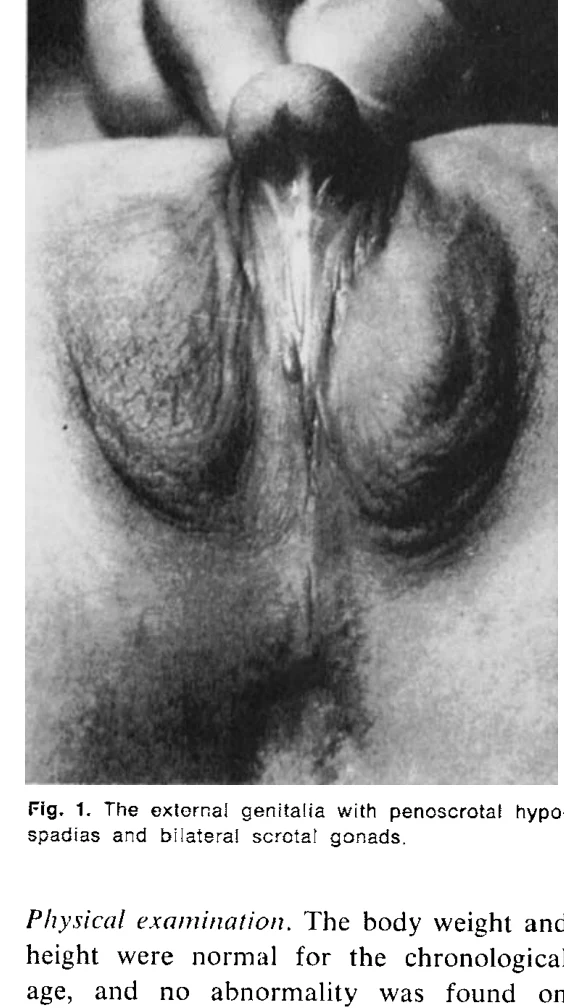
18 months · Infantile 46,XX male DSD (formerly XX male syndrome) with penoscrotal hypospadias.
Browse cases and figures from the original collection. Search by condition, gene, karyotype or phenotype and follow the source publication.
280 Image cases571 Images
Current selection: 280 Image cases · 571 Images
Select an image to enlarge it. Original images retain the complete source file.
1 Images18 months · Infantile 46,XX male DSD (formerly XX male syndrome) with penoscrotal hypospadias.
 1 Images
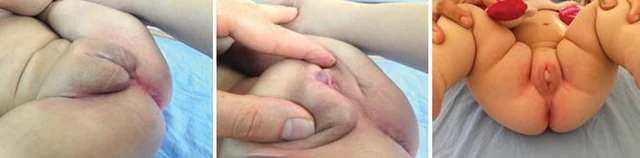
1 year (raised male) · 46,XX male (SRY translocation) with Sertoli cell-only syndrome (SCOS), presenting with ambiguous genitalia.
 6 Images
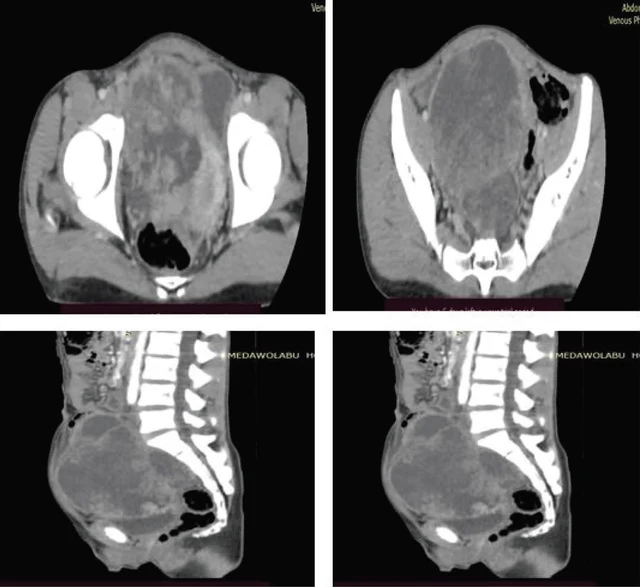
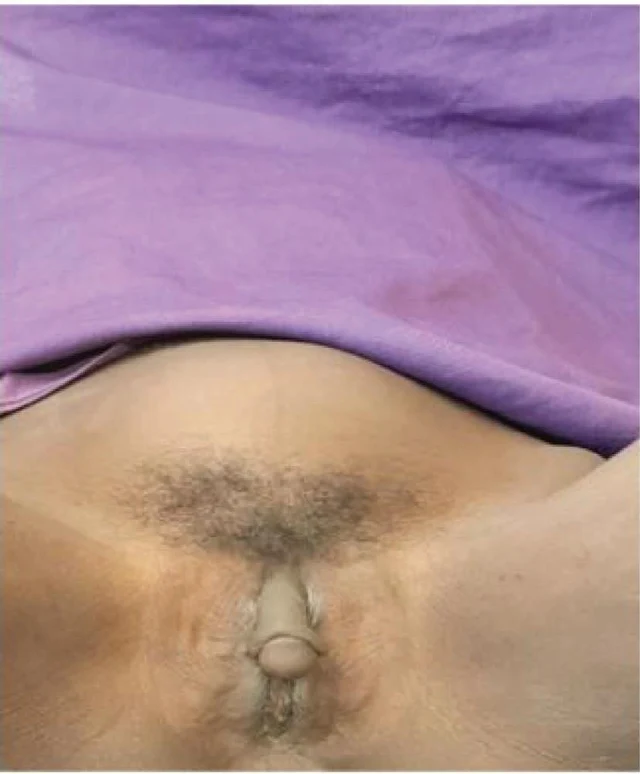
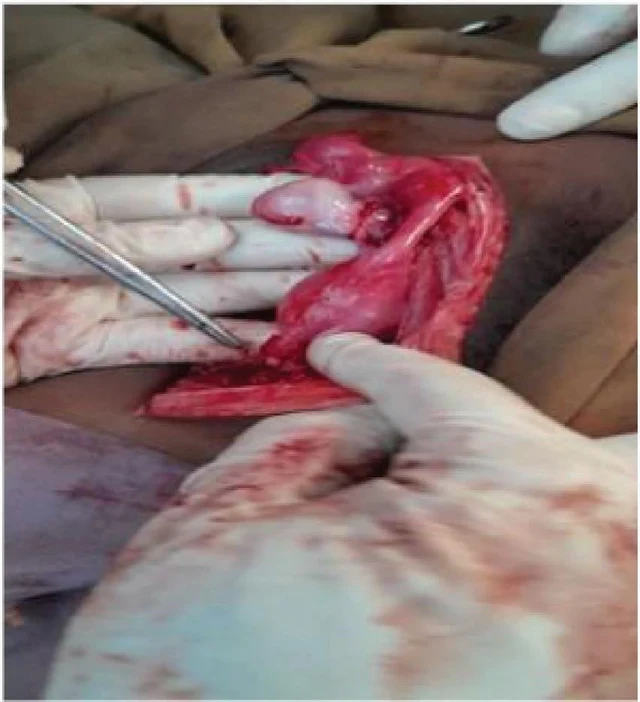
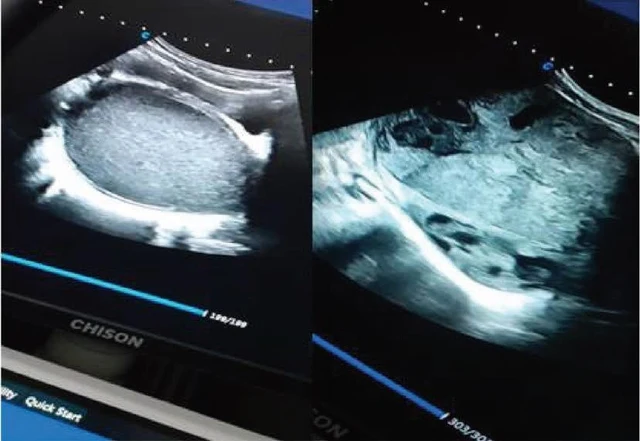
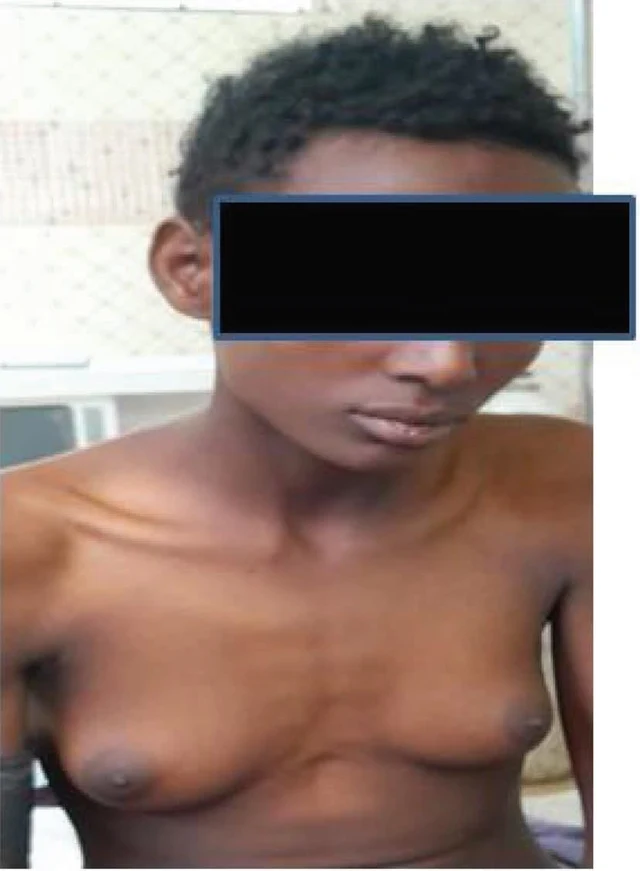
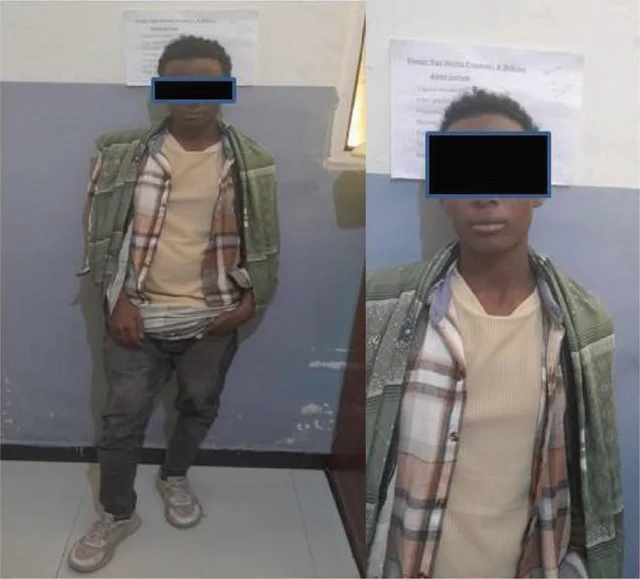
Right-sided tumor pathologically confirmed as dysgerminoma; left-sided mass highly suggestive of synchronous/metachronous dysgerminoma. No ovarian or testicular tissue was found in the sections, so OT-DSD remains only the authors' clinical suspicion.
 1 Images
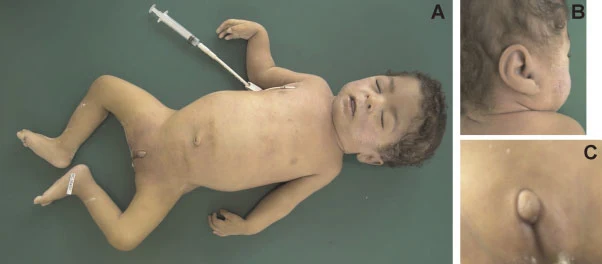
11 months · Partial 16q trisomy (derivative chromosome 15 SMC containing 16q material) with 46,XY DSD and ambiguous genitalia.
 1 Images
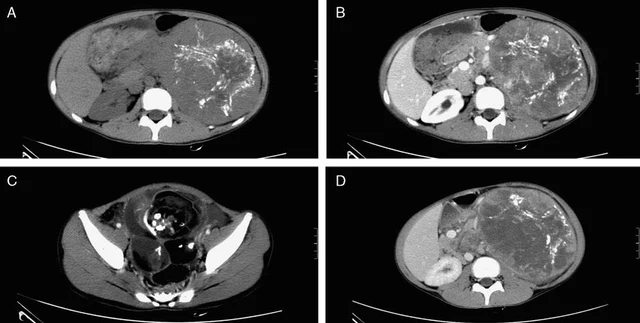
46,XX DSD (androgen excess): female pseudohermaphroditism due to adrenocortical carcinoma and ovarian teratoma.
 1 Images
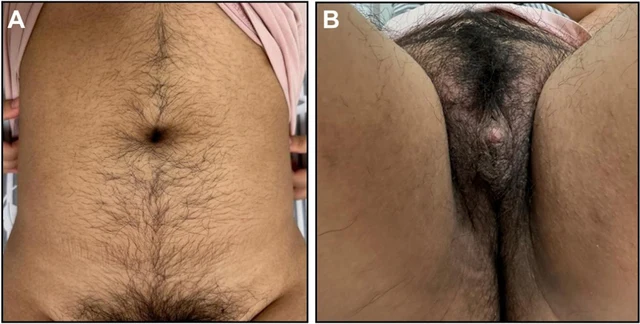
Non-classical congenital adrenal hyperplasia (NCCAH, CYP21A2 deficiency), 46,XX DSD with clitoromegaly/hirsutism.
 3 Images
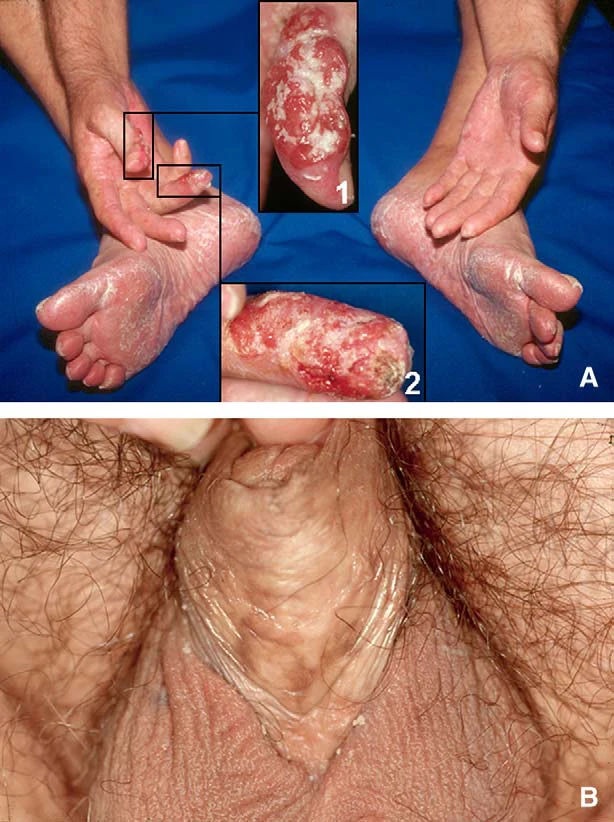
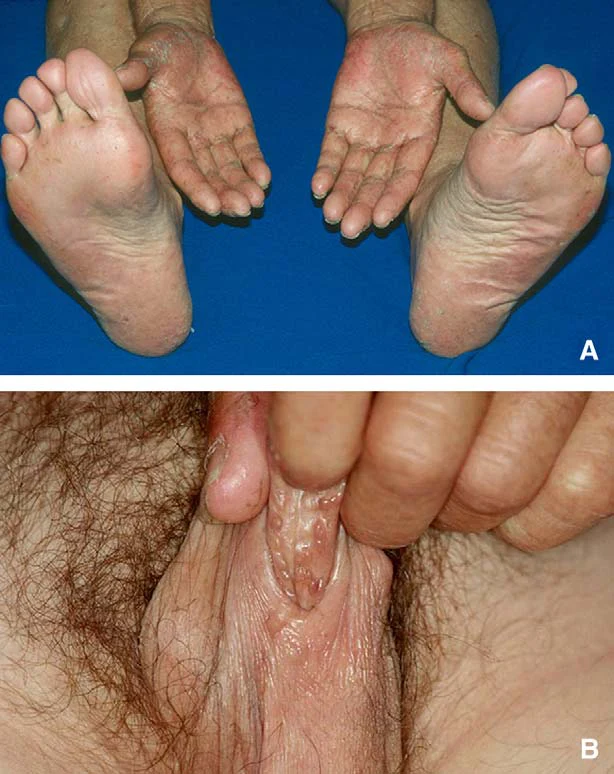
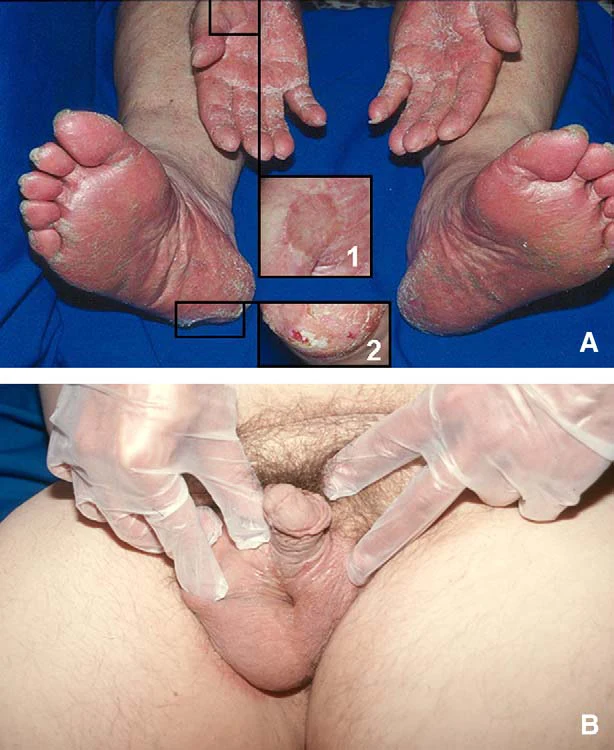
Novel syndrome: palmoplantar keratoderma + cutaneous SCC + dental anomalies + 46,XX gonadal dysgenesis/ambiguous genitalia (hypogenitalism).
 4 Images
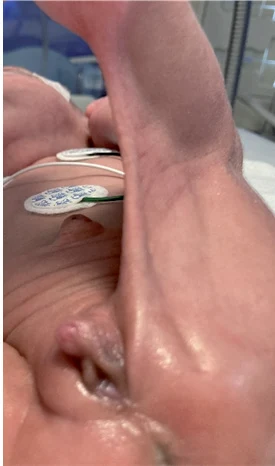
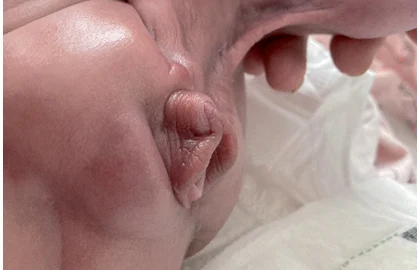
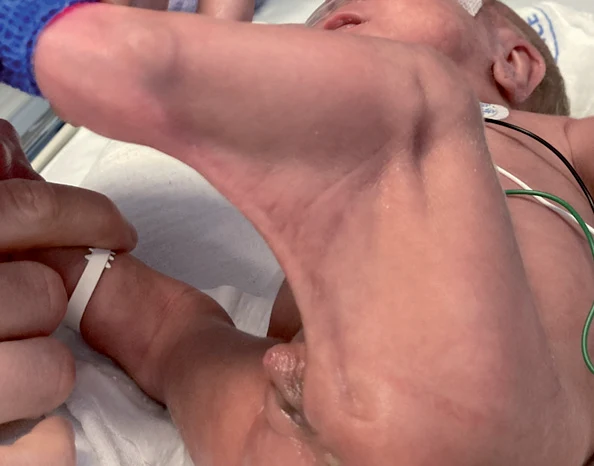
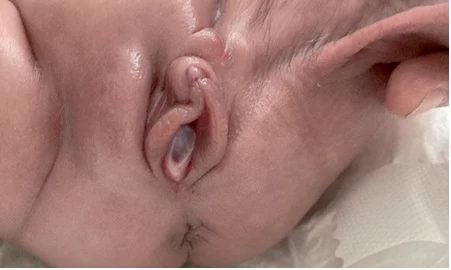
Transient clitoromegaly in a very preterm infant (attributed to preterm-related transient androgen/gonadotropin elevation), non-persistent DSD.
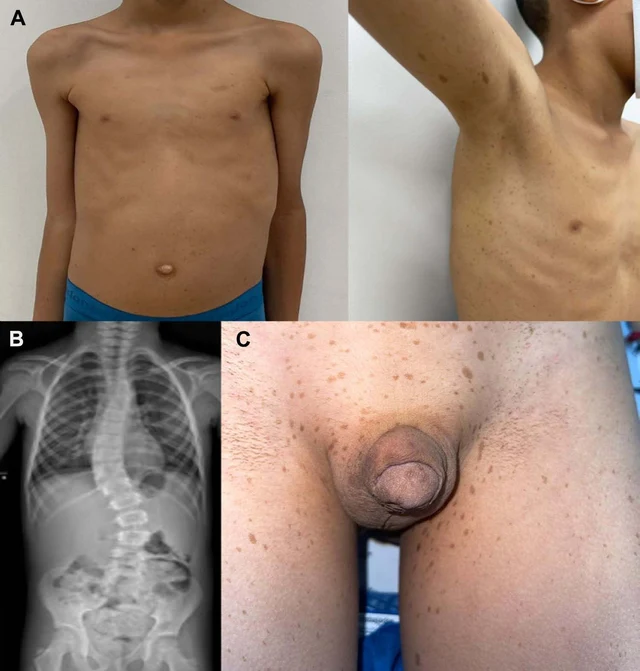 1 Images
46,XY DSD: NF1 combined with NR5A1 variant causing hypospadias/cryptorchidism.
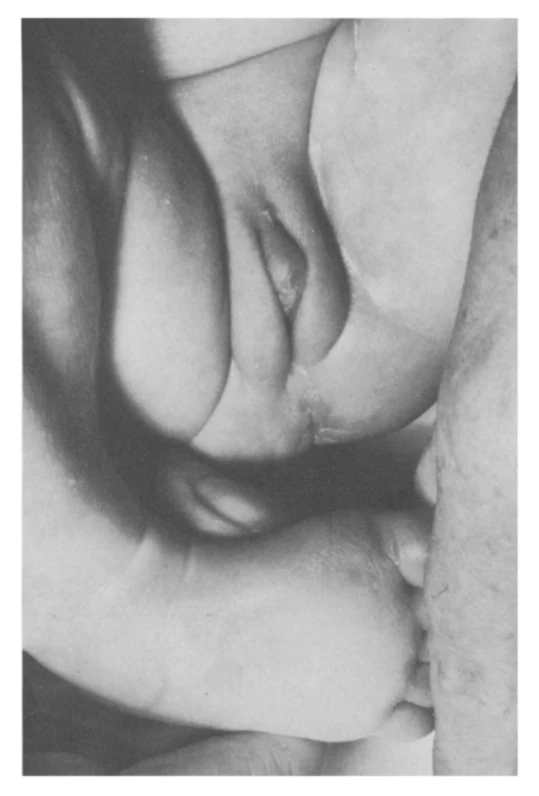 5 Images
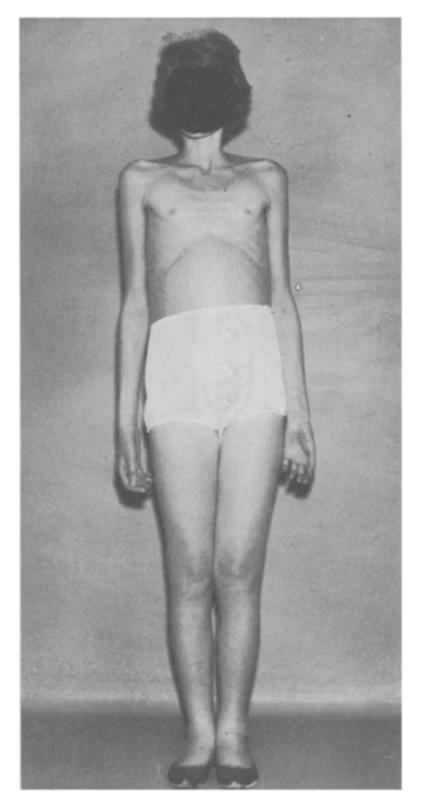
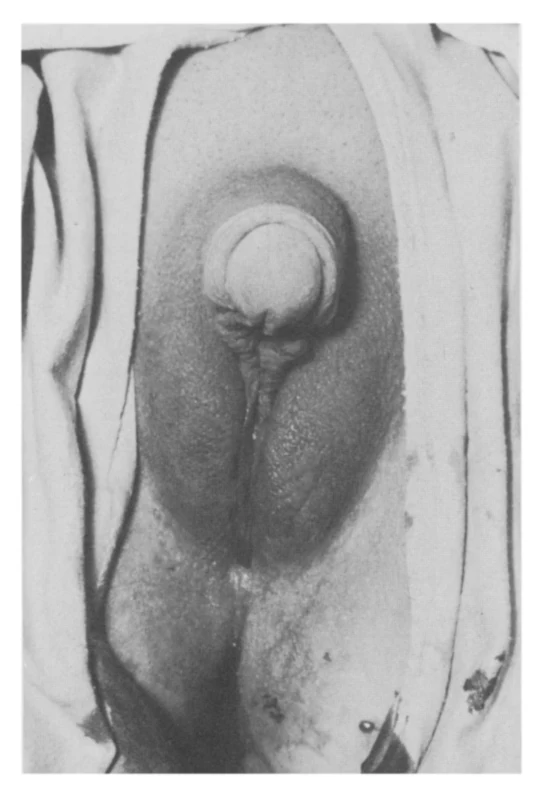
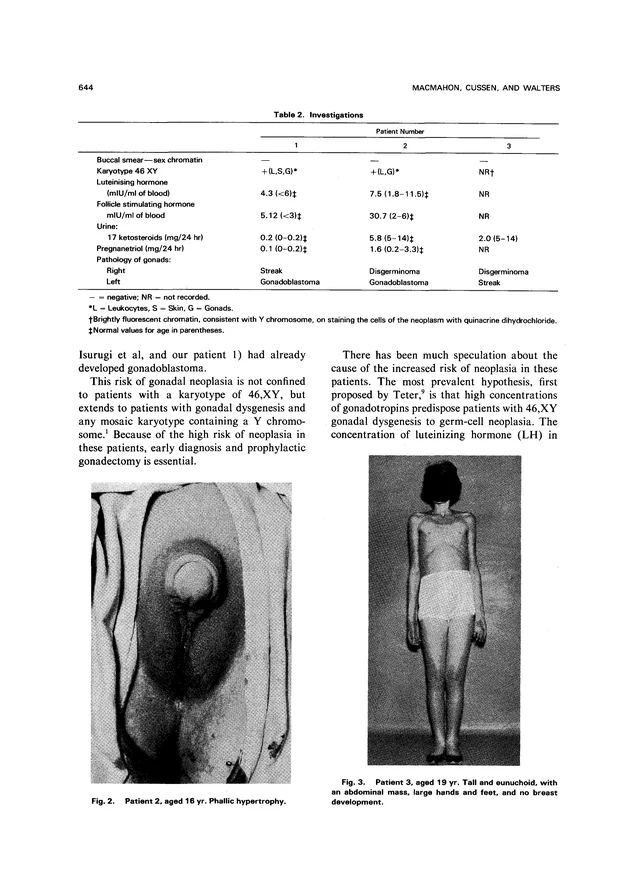
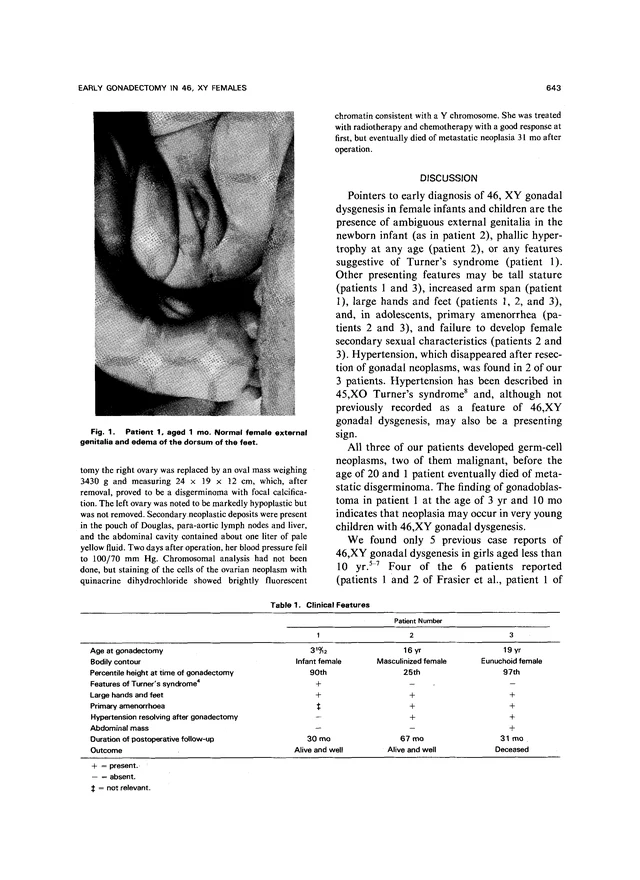
46,XY complete gonadal dysgenesis (Swyer syndrome), all three cases with gonadal tumors/dysgenesis.
 3 Images
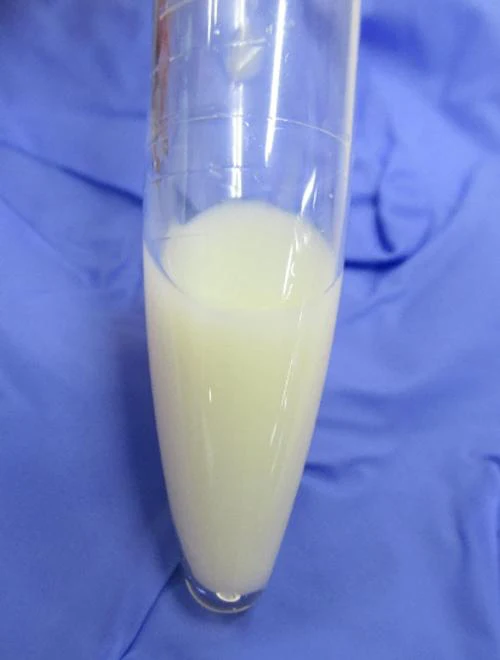
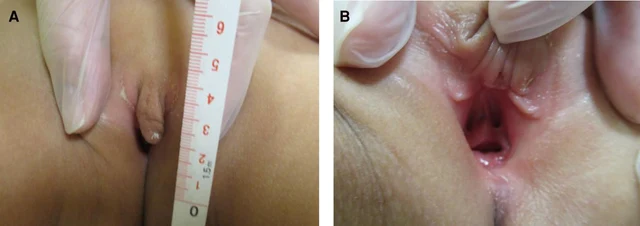
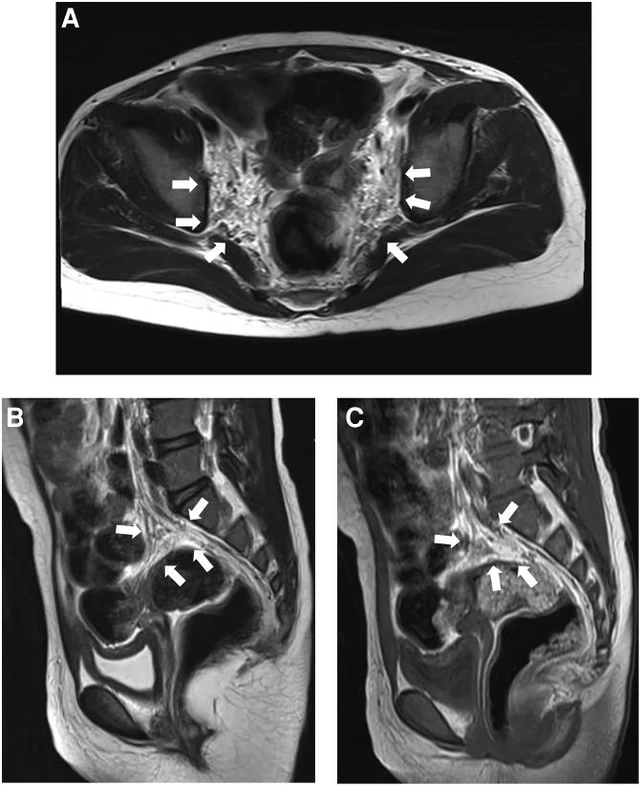
Noonan syndrome (RIT1) with ambiguous genitalia/clitoromegaly and pelvic lymphatic malformation.
 1 Images
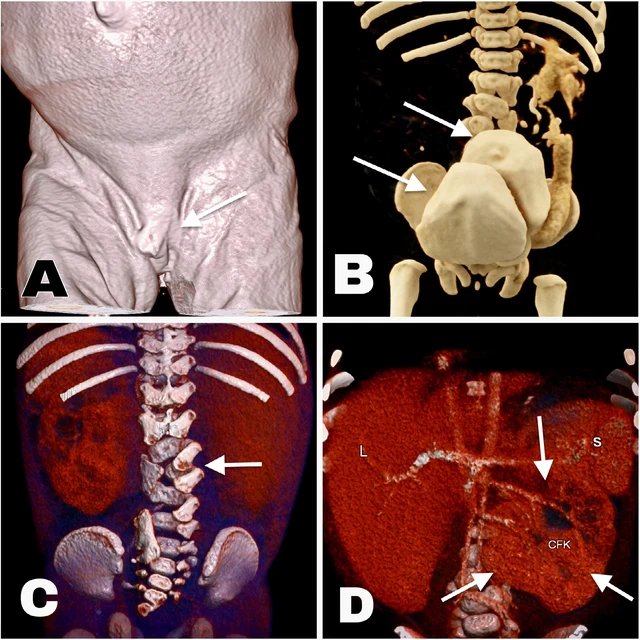
DSD (virilized external genitalia + female chromosomal sex) with duplicated bladder and cross-fused pancake kidney.
 2 Images
Maternal pregnancy luteoma causing virilization of a female fetus (46,XX DSD, androgen excess).
 1 Images
Complete androgen insensitivity syndrome (CAIS), 46,XY male pseudohermaphroditism.
 1 Images
Ovarian steroid cell tumor (OSCT) causing virilization, mimicking PCOS in 46,XX DSD (female virilization).
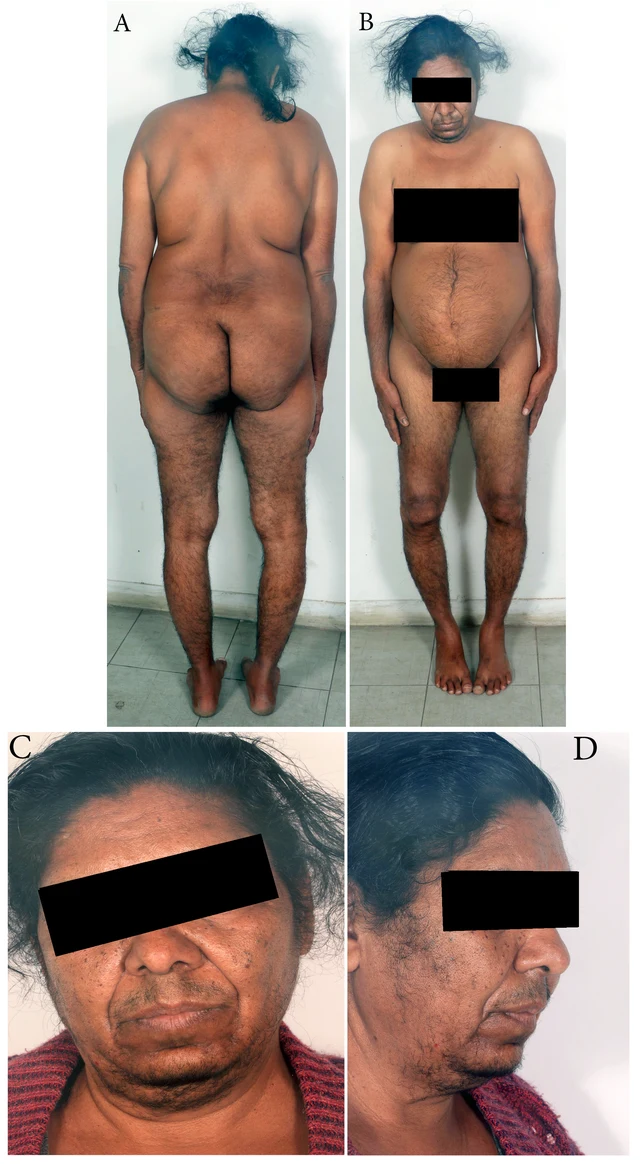 1 Images
GH-ACTH co-secreting pituitary macroadenoma with bilateral functional adrenal adenomas (autonomous cortisol and androgen secretion) causing acquired female virilization (46,XX DSD phenotype).
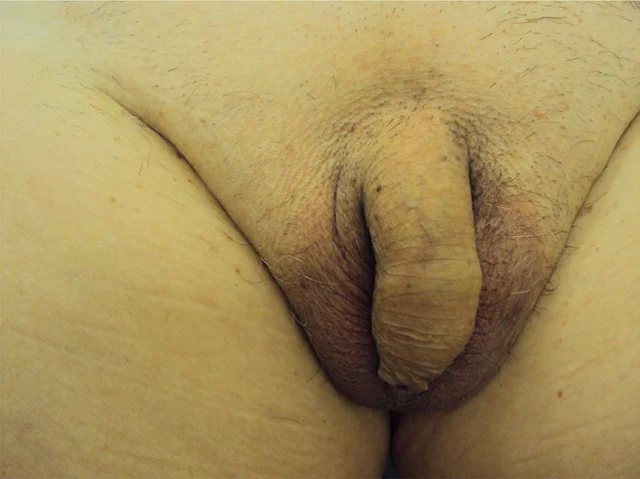 2 Images
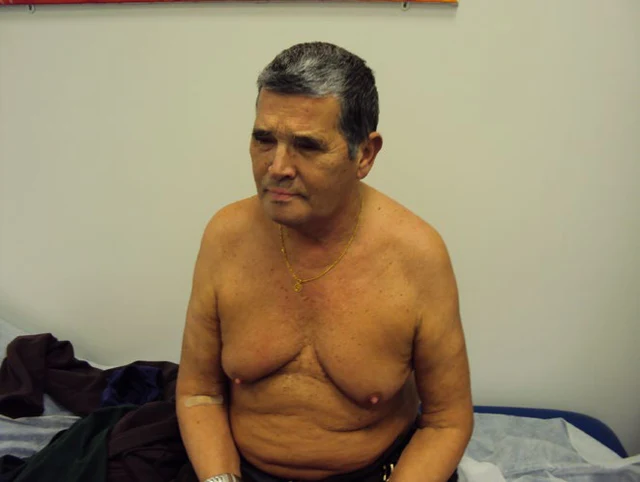
46,XX male / 46,XX sex reversal disorder (SRY-positive), primary hypogonadotropic hypogonadism, secondary non-functioning pituitary macroadenoma causing panhypopituitarism.
 1 Images
Gonadotropin-dependent ovarian theca cell hyperplasia (hyperthecosis) causing postmenopausal virilization (46,XX DSD phenotype).
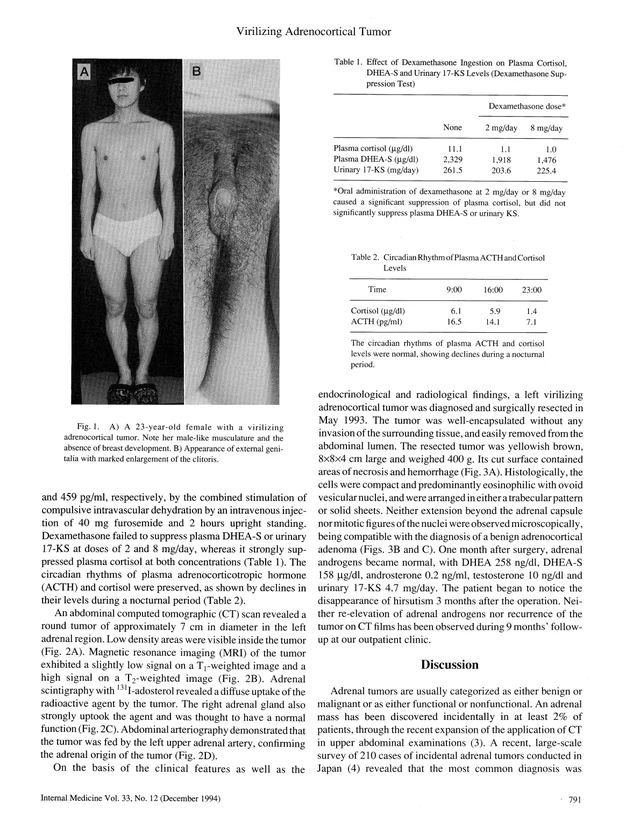 1 Images
Adrenal-genital syndrome due to androgen-secreting adrenocortical tumor (female pseudohermaphroditism / 46,XX DSD)
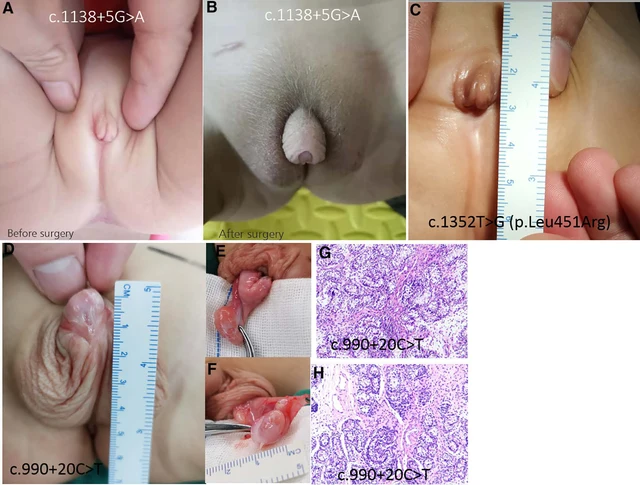 1 Images
NR5A1-related DSD with hypospadias and gonadal dysgenesis (SF-1 deficiency)
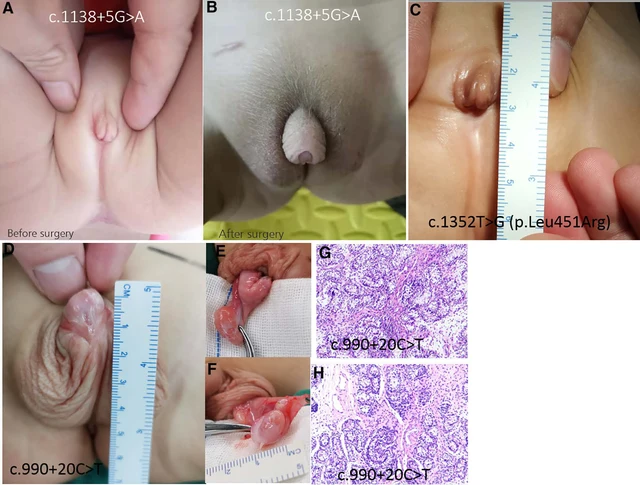 1 Images
Severe NR5A1-related DSD with gonadal dysgenesis (ligand-binding domain variant)
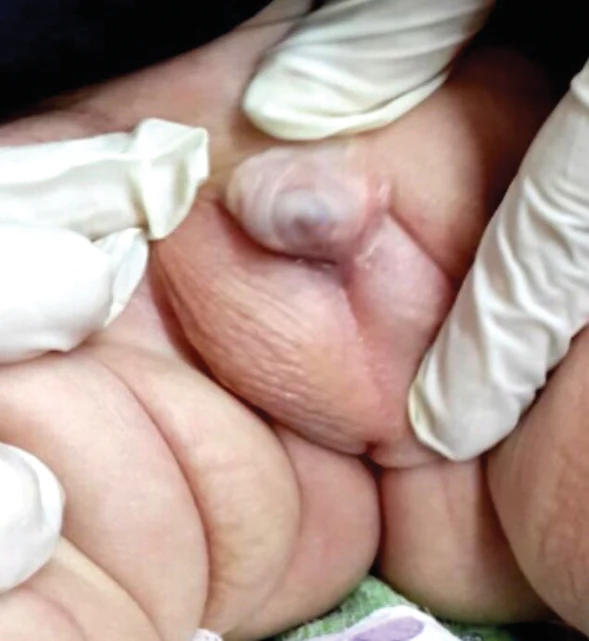 2 Images
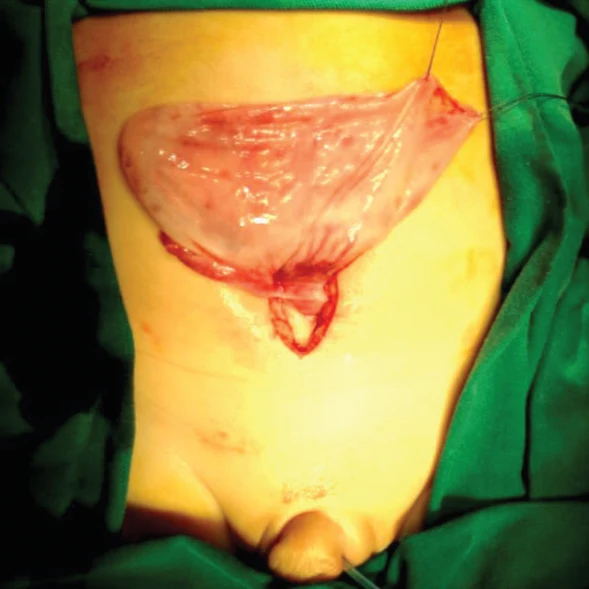
Classic salt-wasting 21-hydroxylase deficiency (congenital adrenal hyperplasia), CYP21A2 R356W homozygous mutation, 46,XX female pseudohermaphroditism (fetal androgen excess causing virilized external genitalia).
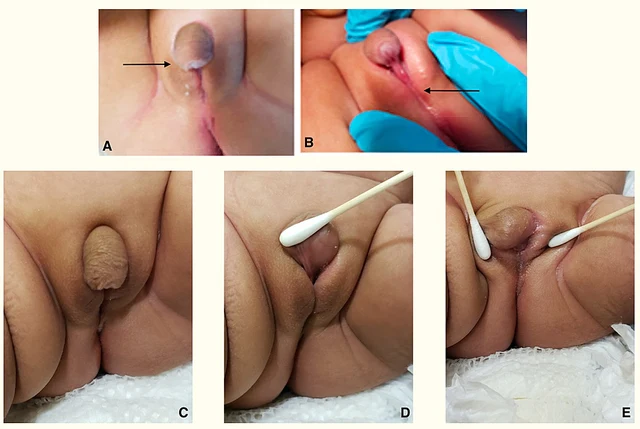 1 Images
Severe penoscrotal hypospadias with sex misassignment in 46,XY DSD (incomplete masculinization / hypospadias spectrum).
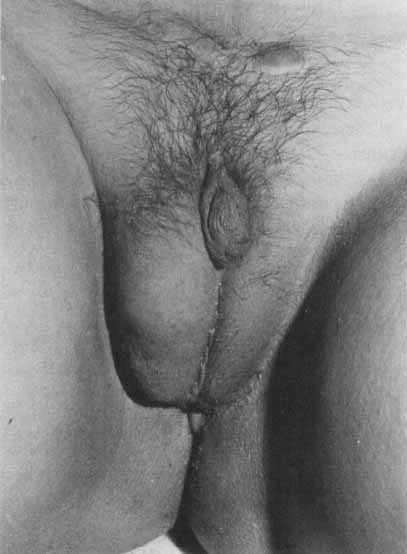 2 Images
Partial androgen insensitivity syndrome (PAIS) / partial androgen resistance - a special subtype with complete absence of androgen receptor in cultured fibroblasts.